
Prof. Dr. Kai S. Exner • Dr. Stéphane Kenmoe
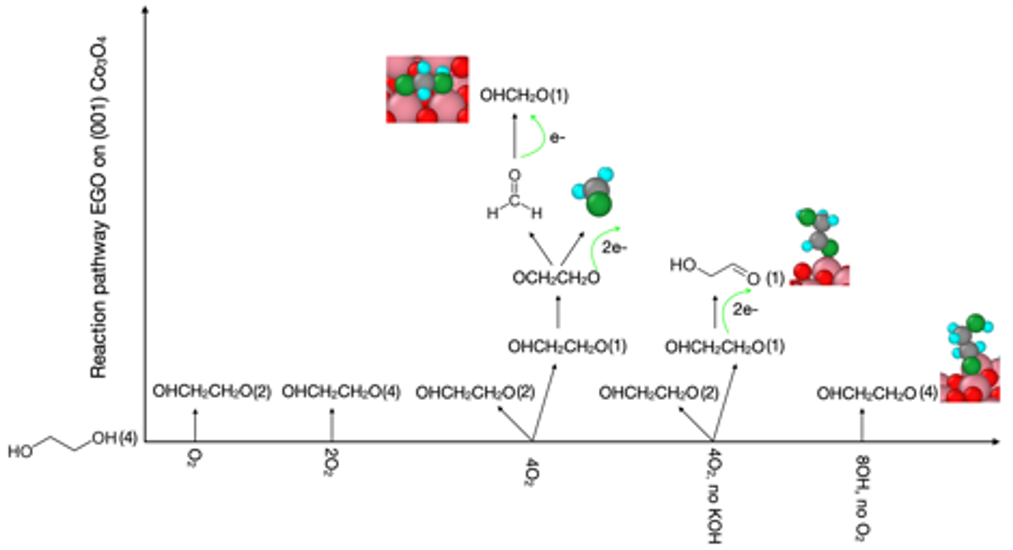
The goal of A06 in the third funding period (FP3) is related to controlling the catalytically active phase to enable selective (electro)-chemical transformations at the solid/liquid interface for both thermal catalysis and electrocatalysis. We are going to continue our work on ethylene glycol oxidation from FP2 by combining insight from the electrode/electrolyte interface (obtained from ab initio molecular dynamics simulations; Kenmoe) with modelling of the elementary steps (through density functional theory calculations; Exner). To this end, we are going to identify optimum conditions for selective ethylene glycol oxidation. For thermal conditions, we scrutinize the impact of different alkali cations (K+, Na+, Cs+) on the frustrated phase transition (restructuring of the catalyst) and catalytic activity, with the aim of correlating the results with temperature, base concentration, and chemical composition of the cobalt oxide surface, considering dopants and vacancies. Related to electrochemistry, we scrutinize the impact of pulsing the electrochemical potential to gain atomic-scale control over the catalytically active phase. It is likely that the catalytically active phase in pulsed experiments is different from the catalytically active phase in constant potential experiments, and this may reduce the degradation and reconstruction of the catalyst under anodic bias. This allows us to exploit different surface phases that potentially circumvent frustrated phase transitions for selective electrochemical transformations.
Figure: Reaction pathway to EG’s thermal and electrocatalytic oxidation intermediates, and products on the Co3O4 (001) surface. The numbers of intermediate and product species present in solution are given in parentheses. Metastable intermediates are not attributed a number. Cobalt atoms are represented in pink, surface oxygen depicted in red, carbon in gray, glycol’s oxygens in green, and glycol’s hydrogens in cyan. [See related publication.]
A06: Density Functional Theory and Molecular Dynamics Studies of Selective Oxidation Reactions of Alcohols and Diols in Aqueous Solutions on Transition Metal Oxides
Thermal and electrochemical oxidation reactions of alcohols on water-covered transition metal surfaces (TMOs) such as Co3O4 and CoFe2O4 spinels can nowadays be described best theoretically by a combination of quantum mechanical density functional theory (DFT), to study stable adsorbate species and transition states, and ab initio molecular dynamics (AIMD) based on DFT, to include ensemble effects and the impact of finite temperature. Ab initio MD studies in this project during the first funding period indicated that the presence of water on TMOs does not only play a significant role in determining surface structure (e.g., by lifting reconstructions), but also produces a large surface structure-dependent number of hydroxyl groups which are instrumental as hydrogen acceptors during electrochemical oxidation of 2-propanol to acetone. Furthermore, the mechanistic details of simulated oxidation reactions vary between those isolated 2-propanol molecules and larger adsorbed clusters of 2-propanol molecules (adsorbed and hydrated), and between different oxidation agents. We performed in-depth studies of the oxidation of 2-propanol to acetone on a hydrogen-deficient hydroxyl group-rich Co3O4 surface covered by a water phase, which may be characteristic for electrochemical oxidation reactions. Contrasting the observed behavior with specific molecular oxidants such as molecular oxygen or adsorbed OOH groups, or even the possibility that oxygen evolution competes with the oxidation of alcohols as evidenced by experimental investigations in projects A01 and A02, opens the exciting perspective to use our theoretical toolbox (in particular by extending DFT-based studies to the ones developed in project B08) to study the selectivity of alcohol oxidation reactions during the second funding period.
Based on the gained experience with 2-propanol oxidation, we plan to extend our scope to the study of different possible reaction channels of this molecule, and continue studying the influence of surface composition, surface defects, and hetero-atom doping, as and when more experimental data become available by a combination of DFT for a screening of mechanistic pathways evaluated by a modified volcano approach, and DFT-based AIMD for solvation and environmental effects. Additionally, we will widen our scope to study ethylene glycol, primarily because of its richer oxidation chemistry, but also because of the industrial importance of its oxidation products.
The long-term focus of this project is on the fundamental mechanistic aspects of alcohol oxidation on TMO surfaces, addressing how the electron-proton transfer steps can be manipulated by suitable environments with the goal to steer selectivity towards desired products.

Copyright © TRR247 2026
Last update: Jul 28, 2026
