
Prof. Dr. Jörg Behler
Project A10 has been established as a new project during the second funding period (FP2) with the aim to gain insights into the catalytic process of alcohol oxidation at Co3O4-water interfaces by performing large-scale reactive molecular dynamics simulations. For this purpose, machine learning potentials (MLP) have been introduced as a new tool in the CRC, which allow to transfer the accuracy of density functional theory (DFT) calculations to simulations beyond the time and length scales accessible by ab initio approaches. We found that MLPs based on optPBE+U DFT reference data can accurately describe the properties of Co3O4 as well as of water. Building on these results, we performed large-scale simulations of the structure, dynamics and reactivity of Co3O4-water interfaces.
In FP3, we will extend this work to characterize the atomistic details of the frustrated phase transition of Co3O4 to CoO under reaction conditions. This dynamic process, which involves disordered and metastable structures including various patterns of oxygen vacancies and cobalt atoms in different oxidation states, is of key importance for the catalytic activity. Due to the complex electronic structure of cobalt-based oxides, non-local fourth-generation high-dimensional neural network potentials (4G-HDNNPs) will be employed. 4G-HDNNPs allow to correctly describe coexisting Co2+ and Co3+ ions, which is a prerequisite for obtaining a reliable potential energy surface for studying complex interface structures and the oxidation reaction itself. To confirm a high-quality description of the electronic and geometric structure of the system, the potential will be validated in collaboration with other theory and experimental groups focusing on the detailed characterization of the catalyst’s surface. In addition, studies of cation doping and anion substitutions will provide further insights into the possibilities to control and modify the catalytic activity by unravelling the interplay between structural, electronic, and catalytic properties.
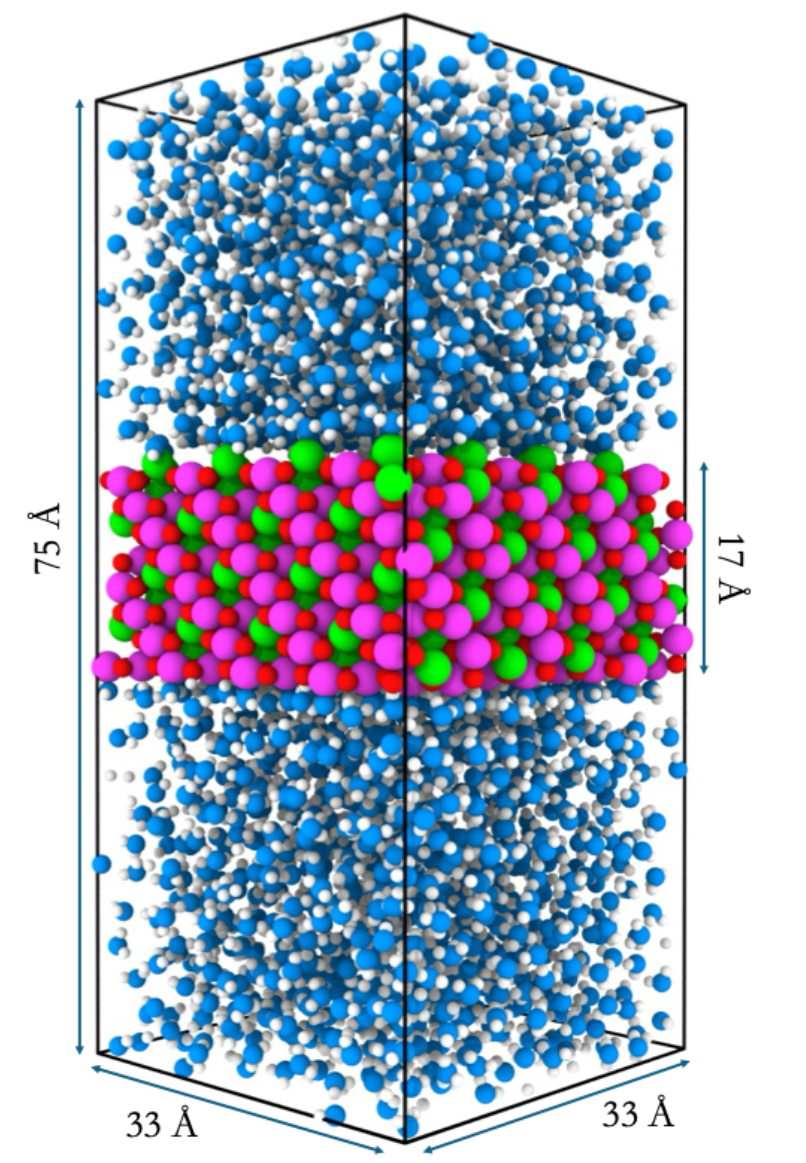
Figure: Slab of the Co3O4(001) surface in contact with water. The slab corresponds to a 4x4x2 Co3O4 supercell and 2048 water molecules resulting in total in 7936 atoms. The top surface of the slab shows the A-termination, while the bottom surface represents the B-termination. Co2+ is shown in green, Co3+ in magenta, oxide oxygen in red, and water oxygen in blue. Figure adapted from related publication.
A10: Molecular Dynamics Simulations of Oxidation Catalysis at Complex Interfaces
Nowadays, computer simulations have become an increasingly important tool, since in principle they allow to gain insights into the atomistic details of catalytic reactions. However, the reliability of the results obtained in these simulations crucially depends on the quality of the underlying potential-energy surface describing the atomic interactions. While the accuracy of first-principles calculations like density functional theory (DFT) is required for reliable and predictive simulations, the time and length scales of DFT-based ab initio molecular dynamics simulations are restricted by the high computational costs, which pose severe limitations on the complexity of the systems that can be studied.
To overcome this limitation, in recent years a lot of effort has been put in the development of machine learning potentials, which allow to combine the accuracy of electronic structure calculations with the efficiency of empirical potentials making them an important new tool in chemistry and materials science. Specifically, in the present project high-dimensional neural network potentials (HDNNPs), which we have developed and applied to many systems in the past one and a half decades, will be used to study the thermal oxidation of small alcohols in aqueous solution at Co3O4 surfaces, because this material has been identified as a very promising catalyst in the first funding period of the CRC.
Our aim is to first address the structural and dynamical properties of a variety of interfaces between Co3O4 and water including a wide range of facets, terminations, defects and surface morphologies as well as the possible formation of surface hydroxides. For this purpose, we will use molecular dynamics (MD) and metadynamics simulations based on HDNNPs trained to DFT+U data, which will allow to significantly extend the time and length scales to nanoseconds and tens of thousands of atoms. Once important structural interface features have been identified, their role as possible active sites will be assessed by studying their interaction and reactivity with 2-propanol in aqueous solution under different conditions like temperature and concentration in close collaboration with other theoretical and experimental groups in the CRC.
In the long-term perspective, in the third funding period the composition of the catalyst might be extended to doped cobalt spinel oxides, to gain insight into the relation between global as well as local composition and catalytic activity. Moreover, our established simulation setup will be applied to the oxidation of further alcohols like ethylene glycol to cover a broader range of systems included in the Comparative Study of the CRC.

Copyright © TRR247 2026
Last update: Aug 17, 2026
